my jobs
login
Register
Welcome back
- Remember me Forgot password?
- Home
- Analysis Platform

Adapter Trimming and Quality Trimming

Quality Control

Taxonomic profiles

Functional profiling

Contig binning

Assess assembly quality

Coverage compute

Binning algorithm

Binning evaluate

de-replicate

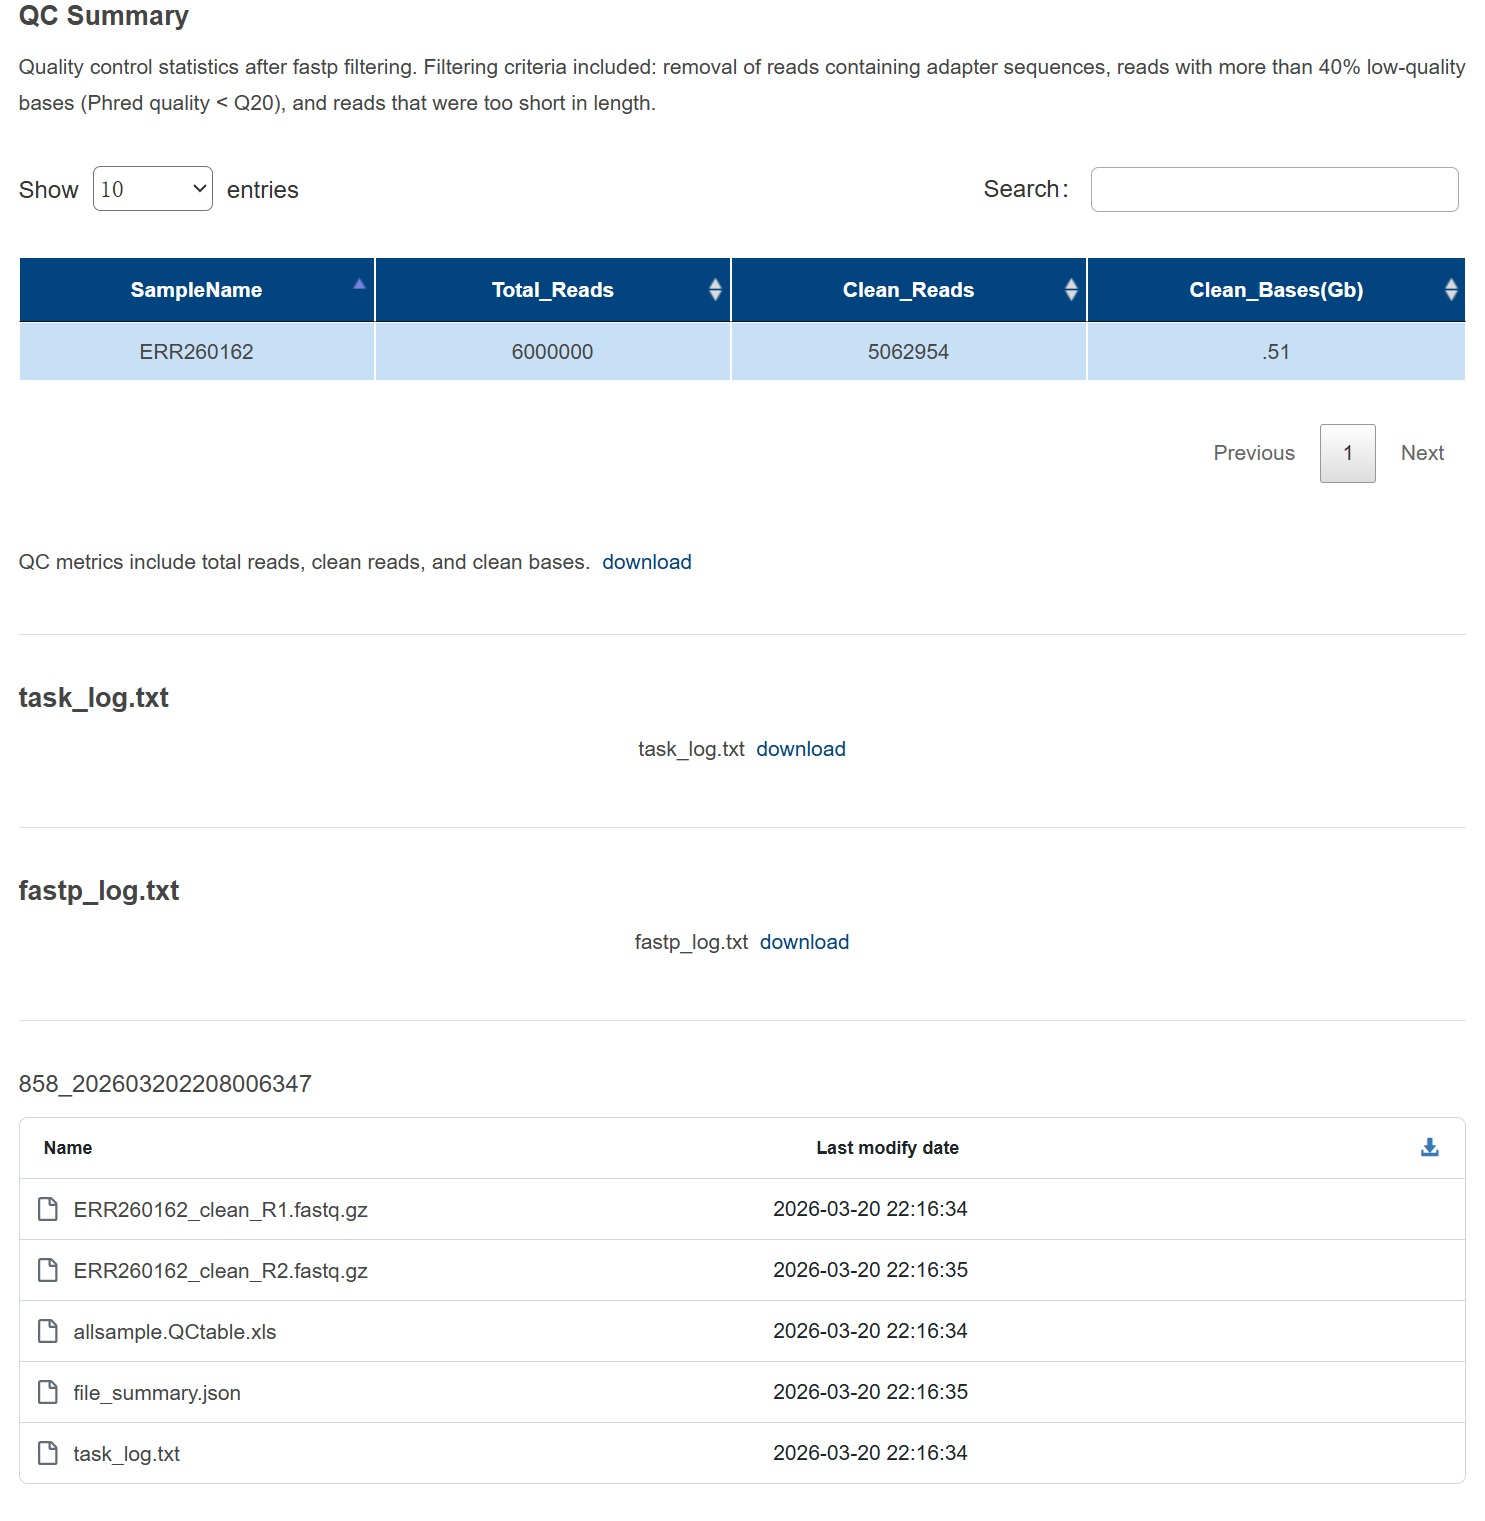
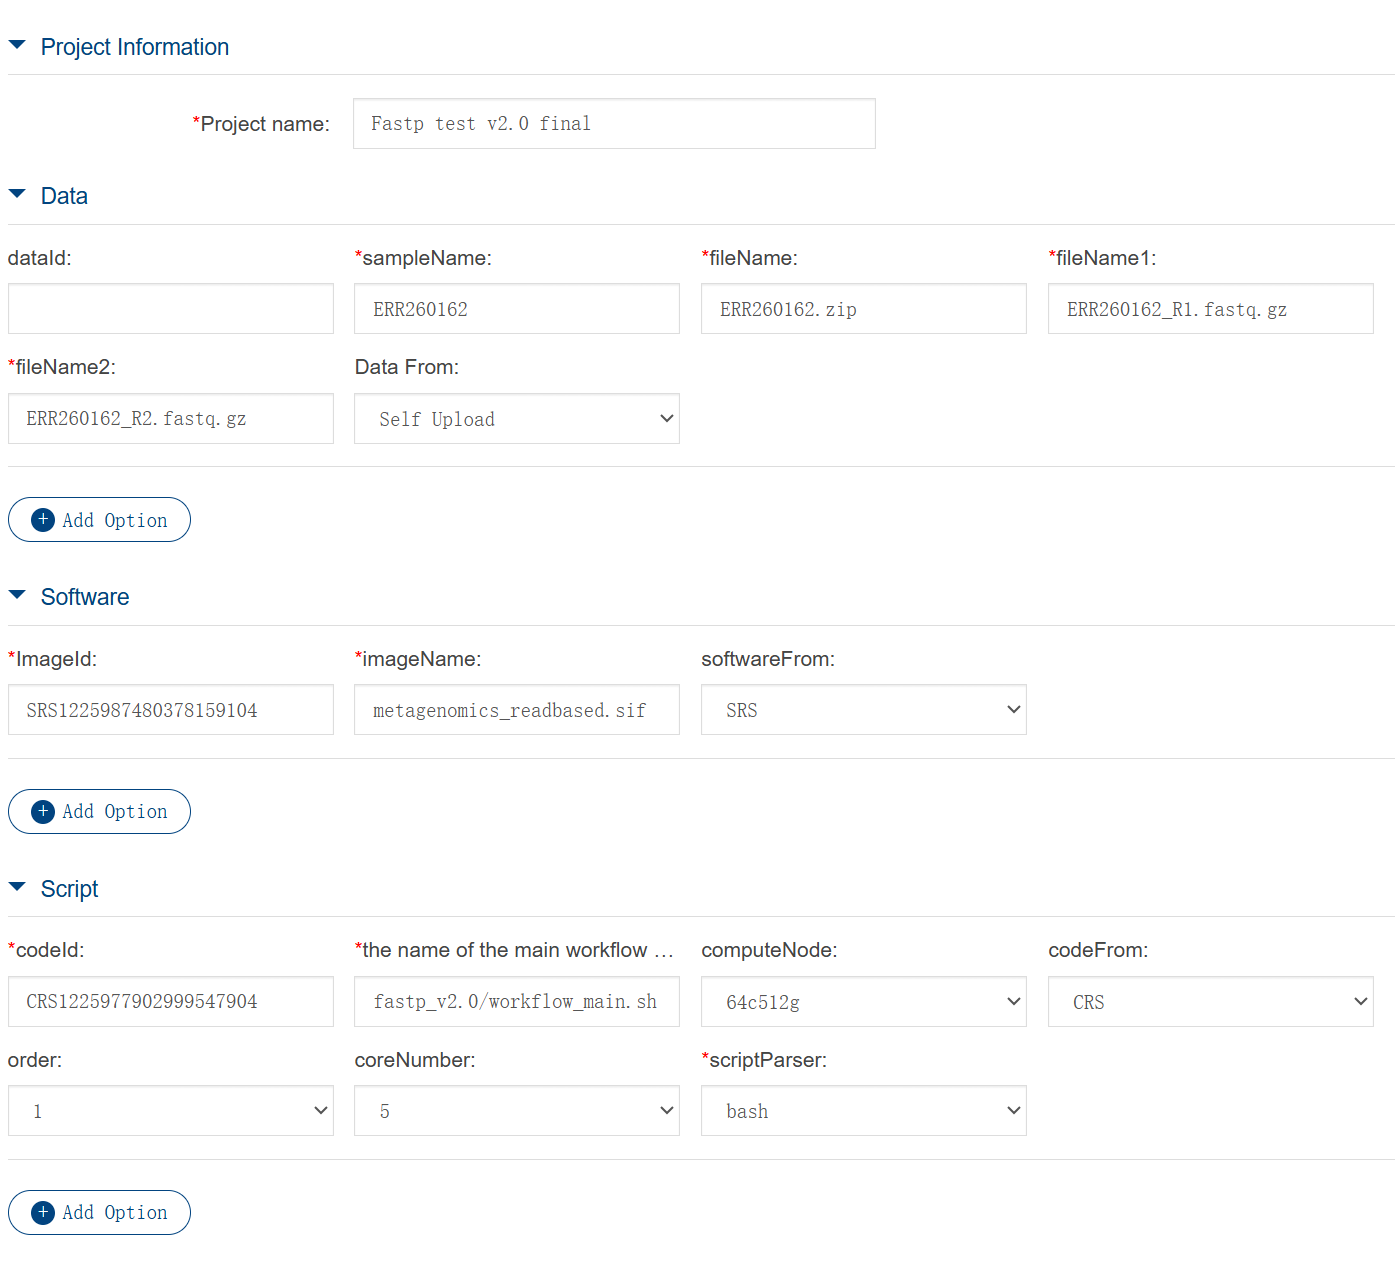
Fastp QC Analysis

GTDB-Tk annotation

Bowtie2 filtering human

SOAPdenovo

CD-Hit

QUAST

vegan

StrainPhlAn

ESMfold

Prodigal ORF analysis

fastqc

Barrnap

tRNAscan-SE Bacterial Analysis

MAFFT

MMseqs2 ORF Clustering

Mash analysis

bcftools

bedtools intersect

Cancer Genomic Analysis

Precise medication guidance

Transcriptome Expression Data Analysis

RNA-seq Pipeline Accuracy Test

Single Cell RNA-seq Analysis

DNA Methylation Sequencing Data Analysis

WGBS Pipeline Accuracy Evaluation

Proteomics analysis platform

RNA-protein binding sites prediction
- Code Repository
- Software Repository
- Data Repository
- Help
- About Us
- News


 Address: 800 Dong Chuan RD. Minhang District, Shanghai, China SJTU-Yale Joint Center for Biostatistics, SJTU